I really want to use some but I’ve heard a lot of skepticism directed towards viome
My GP recommended American gut project. I never tried though, but plan to do soon.
1 Like
Recent articles:
and
Peanut Study: The work was supported by The Peanut Institute- concerning.
1 Like
I used this one:
They check your body’s response to glucose, clearing fat from your blood, and your gut micro-biome.
They then recommend foods for you, food combinations, and meal combinations that work best with your specific makeup.
This helped me make changes with the foods I eat, adding fermented foods, and being more careful of carbs. These changes have been lasting, so I continue to be grateful for the info.
The app has continuing data but I don’t use it any more.
On the gut microbiome front, they list everything they find, but specifically note the 15 gut bacteria that most correlate with ill health and the 15 that most correlate with good health. I was horrified to find I had something like 11 of the bad 15 and only 1 of the good 15. Plus they rate foods on a scale of 1 to 100 and I was eating some things that rated like 6, 12, 9. Thus the diet changes that continue to this day.
Finally, I found it refreshing that foods rating 70 or more can be eaten in any quantity. Their proposition is that calorie counting is less important than overall food quality and nutritional value. I’m not insane so I don’t think I can eat 5 lbs of broccoli for 7 weeks straight and be healthy, but I do like that I can stop counting every last almond and I can eat a few apples if that makes me happy.
2 Likes
1 Like
Genova Diagnostics is comprehensive and used in real clinical situations
Thorne has a new one that looks interesting
2 Likes
deviation from good bugs to bad is accompanied by an increase in gut leakiness, the spillage of bacterial toxins into the bloodstream, and a cascade of inflammatory effects. This has led to the proposal that this microbiome shift is a “primary cause of aging-associated pathologies and consequent premature death of elderly people.”7318
As profound a change in microbiome composition from early adulthood into old age, there’s an even bigger divergence between the elderly and centenarians.7319 When researchers analyzed centenarian poop, they found a maintenance of short-chain fatty acid production from fiber fermentation.7320 For example, in the Bama County longevity region in the Guangxi province of China, fecal sample analyses found that centenarians were churning out more than twice as much butyrate as those in their eighties or nineties living in the same region. If you recall, butyrate is an anti-inflammatory short-chain fatty acid critical for the maintenance of gut barrier integrity. At the same time, there were significantly fewer products of putrefaction, such as ammonia and uremic toxins like p-cresol. The researchers concluded that an increase of dietary fiber intake may therefore be a path toward longevity.7321 An abundance of fiber feeders also distinguished healthy individuals ninety years and older from unhealthy nonagenarians.7322
3 Likes
semi-supercentenarians (those aged 105 to 109) found higher levels of health-associated bacteria, such as Bifidobacteria and Akkermansia.7325 In vaginally delivered, breastfed infants, Bifidobacteria make up 90 percent of colon bacteria, but the level may slip down to less than 5 percent in adult colons and even less in the elderly and those with inflammatory bowel disease.7326 But centenarians carry more of the good bacteria in their gut.7327
Bifidobacteria are often used as probiotics, but anti-aging properties may exist in their postbiotics. Bifidobacteria are one of the many bacteria that secrete “exopolysaccharides,” a science-y word for slime.7328 That’s what dental plaque is, the biofilm created by bacteria on our teeth.7329 Exopolysaccharides produced from a strain of Bifidobacteria isolated from centenarian poop were found to have anti-aging properties in mice, reducing the accumulation of age pigment in their brains and boosting the antioxidant capacity of their blood and livers.7330
Akkermansia muciniphila is named after the late Dutch microbiologist Antoon Akkermans7331 and from Latin and Greek for “mucus-lover.” The species is the dominant colonizer of the protective mucus layer in our gut that is secreted by our intestinal lining.7332 Unfortunately, that mucus layer thins as we age,7333 a problem exacerbated by low-fiber diets. When we eat a fiber-depleted diet, we starve our microbial selves. Our famished flora, the microbes in our gut, have to then compete for limited resources and may consume our own mucus barrier as an alternative energy source, thereby undermining our defenses.7334,7335 Mucus erosion from bacterial overgrazing can be switched on and off on a day-to-day basis in mice supplanted with human microbiomes with fiber-rich and fiber-free diets.7336 You can even show it in a petri dish. Researchers successfully re-created layers of human intestinal cells and showed that dripping fiber (from plantains and broccoli) onto the cells at dietary doses could “markedly reduce” the number of E. coli bacteria breaching the barrier.7337 Aside from eating fiber-rich foods, A. muciniphila helps to directly restore the protective layer by stimulating mucus secretion.7338
A. muciniphila is a likely candidate for a healthy aging biomarker,7339 as its abundance is enriched in centenarians7340 and it is particularly scarce in elders suffering from frailty.7341 A comparative study was undertaken of the microbiomes of people in their seventies and eighties experiencing “healthy” versus “non-healthy” aging, defined as the absence or presence of cancer, diabetes, or heart, lung, or brain disease. Akkermansia, the species most associated with healthier aging, were three times more abundant in the fecal samples of the healthy versus non-healthy aging cohort. Among centenarians, a drop in A. muciniphila is one of the microbiome changes that seems to occur about seven months before death, despite no apparent changes in the physical status, food intake, or appetite at the time.7342 To prove a causal role in aging, researchers showed that feeding A. muciniphila to aging-accelerated mice significantly extended their lifespans.7343
4 Likes
FOOD ADDITIVES TO AVOID
The ultraprocessed foods that make up the majority of our diet7403 aren’t just deficient in fiber but include additives that have been shown to muck with our microbes. Even something as simple as salt can affect our microbiome. Approximately doubling sodium intake by adding a teaspoon of salt to people’s diets not only increases their blood pressure and boosts pro-inflammatory cells7404 implicated in autoimmune disease7405 but it rapidly depletes the gut of the good bacteria Lactobacillus. Nine out of ten study subjects who started out with Lactobacillus in their gut had it completely wiped out by the added salt within just two weeks.7406
2 Likes
Hmm I did send a sample last week after a lot of beets and oat fiber. I did get a lot of theane from a4m in January
| Relative Abundance | Kingdom | Phylum | Class | Order | Family | Genus |
|---|---|---|---|---|---|---|
| 0.1459 | Bacteria | Firmicutes | Bacilli | |||
| 0.1322 | Bacteria | Actinobacteria | Actinobacteria | Actinomycetales | Corynebacteriaceae | Corynebacterium |
| 0.1148 | Bacteria | Cyanobacteria | Chloroplast | Streptophyta | ||
| 0.0980 | Bacteria | Actinobacteria | Actinobacteria | Actinomycetales | Corynebacteriaceae | Corynebacterium |
| 0.0573 | Bacteria | Firmicutes | Bacilli | Bacillales | Bacillaceae | Bacillus |
| 0.0454 | Bacteria | Actinobacteria | Actinobacteria | Actinomycetales | Propionibacteriaceae | Propionibacterium |
| 0.0367 | Bacteria | Proteobacteria | Gammaproteobacteria | Pseudomonadales | Pseudomonadaceae | Pseudomonas |
| 0.0288 | Bacteria | Firmicutes | Clostridia | Clostridiales | [Tissierellaceae] | Finegoldia |
| 0.0277 | Bacteria | Firmicutes | Bacilli | Bacillales | Bacillaceae | Bacillus |
| 0.0273 | Bacteria | Actinobacteria | Actinobacteria | Actinomycetales | Corynebacteriaceae | Corynebacterium |
| 0.0227 | Bacteria | Actinobacteria | Actinobacteria | Actinomycetales | Corynebacteriaceae | Corynebacterium |
| 0.0195 | Bacteria | Cyanobacteria | Chloroplast | Streptophyta | ||
| 0.0191 | Bacteria | Bacteroidetes | Bacteroidia | Bacteroidales | Prevotellaceae | Prevotella |
| 0.0181 | Bacteria | Firmicutes | Bacilli | Lactobacillales | Streptococcaceae | Streptococcus |
| 0.0099 | Bacteria | Firmicutes | Clostridia | Clostridiales | [Tissierellaceae] | Anaerococcus |
| 0.0092 | Bacteria | Proteobacteria | Gammaproteobacteria | Pseudomonadales | Pseudomonadaceae | Pseudomonas |
| 0.0087 | Bacteria | Cyanobacteria | Chloroplast | Streptophyta | ||
| 0.0081 | Bacteria | Actinobacteria | Actinobacteria | Actinomycetales | Corynebacteriaceae | Corynebacterium |
| 0.0071 | Bacteria | Proteobacteria | Alphaproteobacteria | Sphingomonadales | Sphingomonadaceae | |
| 0.0066 | Bacteria | Bacteroidetes | Flavobacteriia | Flavobacteriales | [Weeksellaceae] | Chryseobacterium |
| 0.0061 | Bacteria | Firmicutes | Clostridia | Clostridiales | [Tissierellaceae] | Anaerococcus |
| 0.0055 | Bacteria | Actinobacteria | Actinobacteria | Actinomycetales | Actinomycetaceae | Actinomyces |
| 0.0048 | Bacteria | Cyanobacteria | Chloroplast | Streptophyta | ||
| 0.0044 | Bacteria | Actinobacteria | Actinobacteria | Actinomycetales | Actinomycetaceae | Actinomyces |
| 0.0044 | Bacteria | Proteobacteria | Gammaproteobacteria | Pseudomonadales | Moraxellaceae | Acinetobacter |
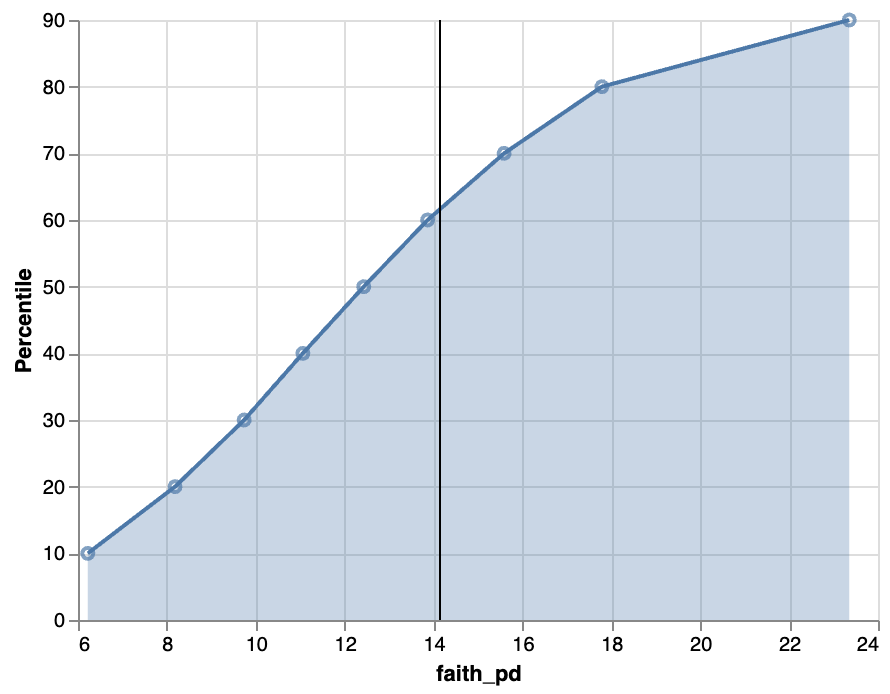
Microsetta - Sample Report (2).csv (5.6 KB)
Microsetta - Sample Report (1).csv (2.6 KB)
Microsetta - Sample Report (8).csv (3.7 KB)
Microsetta - Sample Report (7).csv (13.3 KB)
Microsetta - Sample Report (6).csv (7.6 KB)
historical data i only finally just fed to claude to do cross comparison
1 Like
It looks like the microsetta project has paused for new enrollment. Is there any consensus around another credible microbiome test?
So, bad news is: you’re in the ~20% of people whose sample contains cholesterol, but no coprostanol. This is bad news because the body has a hard time getting rid of cholesterol unless it’s been converted to coprostanol by gut bacteria, so the absence of this microbial function may contribute to high blood cholesterol levels and cardiovascular disease risk.
Coprostanol production appears to be one of the core conserved features of the mammalian gut microbiome—meaning all mammals, including humans, are supposed to have bacteria that do it. However, it seems that antibiotics can drive these bacteria extinct in a person’s gut: in a 1990 study, researchers found that, about 10% of the time, a week-long course of antibiotics caused fecal coprostanol levels to drop to zero—and stay there, even long after the antibiotics had left the system.
new sample!
1. Your microbiome fits your coprostanol-negative result almost suspiciously well
This may be the coolest finding.
Your earlier stool chemistry found cholesterol present but no detectable coprostanol . In the sequencing sheet, several taxa in/near the bacterial lineages now implicated in cholesterol → coprostanol metabolism are completely absent:
- CAG-177: 0
- Fimenecus: 0
- Acutalibacter: 0
- every listed
Eubacterium_*group: 0That is intriguing because the original
ismAwork placed cholesterol-metabolizing organisms in an uncultured Clostridium-cluster-IV-like lineage. Even more interestingly, a 2026 Nature Communications paper identified another enzyme, SpiR , that predicts cholesterol conversion better thanismA;spiRappears concentrated in an uncultured Acutalibacteraceae clade, including cholesterol-reducing CAG-177-related organisms. NatureSo you may have something considerably more specific than generic “low diversity”:
a missing metabolic guild.
That’s precisely the distinction you were talking about earlier between taxonomic diversity and response/function diversity . You can have loads of excellent carbohydrate fermenters and nevertheless have a hole in one particular metabolic module.
Caveat: this spreadsheet is taxonomy, not gene annotation. It cannot prove that
spiRorismAis absent. Shotgun gene-level analysis of your reads is the test I’d really want.
2. You have an enormous mystery organism: CAG-303
This was easy to overlook.
CAG-303 = 3.86% of your entire microbiome.
Only 4/27 people in this batch have any detectable CAG-303 at all, and you’re massively the highest:
- you: 3.86%
- next person: ~0.71%
- next: ~0.59%
- next: ~0.27%
So roughly 1 in every 26 reads assigned at genus level belongs to this poorly characterized lineage.
At species level it is essentially all:
CAG-303 sp000437755
There is frustratingly little mechanistic literature on it. That’s microbial dark matter rather than a familiar probiotic species. A recent metagenomic study happened to find this same genome-defined species enriched in coronary-artery-disease cases, but that is an observational association in a small population and absolutely does not mean your CAG-303 is causing cardiovascular disease. PubMed Central (PMC)
Given how extraordinarily abundant it is in you , I’d put this organism near the top of the list for future functional annotation.
3. You have low global diversity but impressive local microdiversity
This is deliciously paradoxical.
Your overall species count is low, yet your Faecalibacterium population is spread across multiple genomic groups:
- F. prausnitzii_G : 6.24% , highest in cohort
- F. prausnitzii_J : 3.31% , highest
- F. prausnitzii_C : 3.09% , ~top of cohort
- F. prausnitzii_D : 0.90%
Together ≈ 13.5% .
Modern genomics shows that what historically got called F. prausnitzii is actually a highly diverse complex containing multiple deep genomic clades with potentially different functional repertoires. Multiple distinct strains/clades can coexist in one individual. PubMed Central (PMC)
So your microbiome isn’t simply:
few species → little complexity.
It’s more like:
few broad guilds, but some guilds contain several parallel specialists.
That is much more interesting ecologically.
4. Your butyrate-producing network is ridiculous
It’s not just Faecalibacterium.
You are #1 or nearly #1 in the cohort for several organisms capable of participating in butyrate-producing trophic networks:
- Agathobaculum 1.90% , #1
- Butyribacter 2.33% , #1
- Gemmiger formicilis 3.71% , #1–2
- Faecalibacterium 13.54% , #2
- Anaerostipes 4.11% , #2
- Fusicatenibacter 6.77% , #2
- plus Agathobacter, Coprococcus etc.
Agathobaculum butyriciproducens literally produces butyrate as a major fermentation end-product. Gemmiger formicilis was originally described as a carbohydrate-fermenting organism producing both formate and butyrate . PubMed
Your gut therefore seems to have done something like:
compress taxonomic diversity while preserving, or perhaps concentrating, one particular metabolic basin.
That could explain how you can simultaneously have low richness yet seemingly formidable fermentation capacity.
5. Your Bifidobacterium story is unusually concentrated
You have 8.56% Bifidobacterium , roughly 8× the cohort median.
But most of it isn’t distributed across lots of bifidobacterial species. It is overwhelmingly:
B. pseudocatenulatum = 6.62%.
Only 10/27 samples even have detectable B. pseudocatenulatum , and you’re #2.
This one is especially interesting given our diet discussion because some strains possess a specialized xylan-utilization system. Human dietary interventions have shown that long-chain xylan can selectively expand endogenous B. pseudocatenulatum when the relevant xylanase machinery is present. Nature
And you simultaneously have:
Bacteroides xylanisolvens = 1.27% , #2 in the cohort.
That starts looking less like random high fiber intake and more like a fairly coherent xylan-processing consortium .
Potentially:
plant xylan → primary degraders → oligosaccharides/acetate/lactate → secondary fermenters → butyrate
You have organisms sitting at several stations along that conveyor belt.
6. Lachnospira is even stranger at species level than genus level
Your Lachnospira = 9.29% was already #1.
But:
Lachnospira eligens_A = 7.58%.
The cohort median is only 0.20% .
So you’re roughly 37× the median abundance .
And you also have
Lachnospira sp000436475at 1.31% , again the highest person in the batch, with that organism detected in only 3/27 people.That is an extraordinary amount of ecological real estate devoted to a very narrow Lachnospira niche.
7. You’re also #1 for Sutterella
Sutterella = 1.08%
That’s about 4.4× the cohort median and the highest sample.
I wouldn’t label this either good or bad. Sutterella wadsworthensis is a relatively common human gut organism and isn’t generally considered a pathogen, although it interacts with the host/mucosal environment and has produced a zoo of inconsistent disease associations. PubMed Central (PMC)
It matters here mostly because it is another example of your community being extremely non-average in composition despite its low richness.
The pattern I’m starting to see
Your gut doesn’t look like a randomly impoverished microbiome.
It looks more like an ecosystem that underwent a strong ecological bottleneck and then reorganized into several extremely successful functional guilds :
plant-polysaccharide specialists
→ Bifidobacterium / Bacteroides / Lachnospira / KLE1615↓ cross-feeding
SCFA/butyrate network
→ Faecalibacterium / Anaerostipes / Agathobaculum / Gemmiger / Butyribacterwhile apparently losing a bunch of other ecological modules, possibly including the cholesterol→coprostanol module .
That’s much closer to low response diversity with strong redundancy inside selected functions than simply “bad low-diversity microbiome.”
And the craziest individual organism to me isn’t KLE1615 anymore. It’s probably CAG-303 at 3.86% . You have a huge population of something science barely knows what to do with.
The killer next analysis would be to take your raw shotgun reads and map CAZymes + butyrate pathways +
ismA+spiR+ bile-acid genes , then measure functional diversity rather than taxonomy. That could tell us whether your 55-species ecosystem is functionally tiny or whether it has quietly packed a much larger metabolic repertoire into fewer genomes.


Yes, but only at the level of plant classes / fiber chemistry , not exact foods.
The strongest inference from your microbiome is:
- Pectin-rich fruits and vegetables are very likely important. Your huge Lachnospira eligens signal is the clearest clue. L. eligens is a strong pectin utilizer, and pectin is especially abundant in apples, citrus, carrots, beets, sweet potato, and many other fruits/roots . PubMed
- Whole grains / bran, especially wheat, rye, barley, possibly oats/corn are also strongly suggested. Your high B. pseudocatenulatum plus Bacteroides xylanisolvens points toward xylan/arabinoxylan , which is heavily represented in cereal cell walls and bran. In an actual human intervention, wheat-bran arabinoxylan selectively expanded B. pseudocatenulatum strains able to use long-chain xylan. Nature
- Alliums and fructan-rich vegetables are plausible: onion, garlic, leek, asparagus, chicory/Jerusalem artichoke , plus some wheat. This comes partly from your KLE1615 genomic prediction for fructan utilization, but that part is weaker because KLE1615 has not been experimentally characterized nearly as well. Fructans are concentrated in those foods. PubMed Central (PMC)
- Legumes / seeds / some nuts are also plausible from the mannan side, especially soy, beans/other legumes, guar-like seed gums , and potentially coffee-derived mannans. Again, this is a moderate-confidence inference because the KLE1615 β-mannan assignment is computational rather than experimentally proven. ScienceDirect
If I had to reconstruct your diet blind from the stool alone, I’d guess something like:
Lots of fruit/vegetable soluble fiber + substantial cereal/bran fiber + frequent legumes/alliums, rather than mostly leafy greens and salads.
The cereal signal is particularly interesting because your combination of B. pseudocatenulatum + B. xylanisolvens is almost a little xylan neon sign. B. xylanisolvens was isolated specifically for its ability to degrade xylan, and it can attack fibers from wheat/corn bran, peas, cabbage and leeks. PubMed
What I cannot infer well:
Leafy greens versus broccoli versus zucchini versus berries . Those foods certainly affect the microbiome, but your taxa don’t give a clean enough fingerprint to distinguish them.
Likewise, I wouldn’t confidently infer lots of resistant starch foods such as cooled potatoes/rice specifically. Your community can certainly ferment them, but the taxa you have are more loudly shouting pectin + xylan/arabinoxylan + fructan/mannan .
So, ranked by how strongly your microbiome seems to whisper their presence:
1. Whole grains/bran
2. Pectin-rich fruit/root vegetables
3. Onion/garlic/leek/chicory-type plants
4. Legumes / seed-derived fibers
5. Everything else, much harder to distinguishOne funny possibility: your gut may be telling us more about the plant cell walls you eat than about the plants themselves. Two completely different foods can feed the same guild if they deliver the same polysaccharide architecture. That is why microbiome archaeology works better at reconstructing “xylan-rich grasses” than “you ate rye bread Tuesday.”


